Archives: Published Articles

Preparing CDMOs and Sponsors for Regulatory Inspections

For contract development and manufacturing organizations (CDMOs), navigating regulatory inspections can become complex when multiple clients have different specifications, requirements, and expectations. While sponsors remain responsible for ensuring their contract organizations comply with applicable GMP requirements, effective inspection readiness requires close collaboration between both parties.


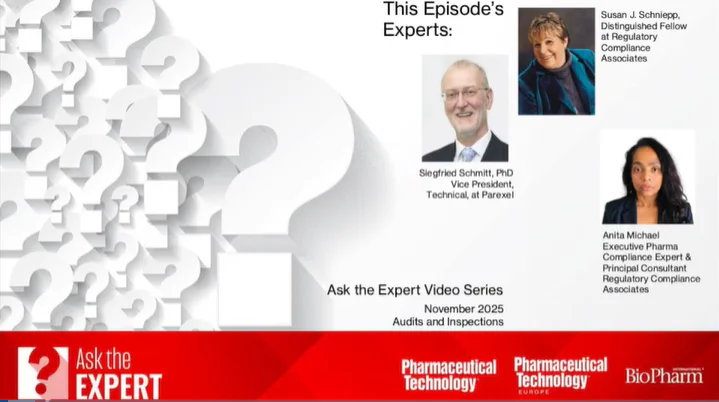
In this Ask the Expert discussion, Susan J. Schniepp and Anita Michael of Regulatory Compliance Associates (RCA), a Nelson Labs company, and Siegfried Schmitt of Parexel discuss how CDMOs and their clients can prepare for inspections and respond to regulatory actions.


The experts emphasize that inspection readiness should extend beyond preparing for an individual client inspection. CDMOs should maintain a robust quality system, supported by effective CAPA and investigation processes, deviation management, KPIs, and holistic tracking and trending. Facilities should also be prepared to demonstrate compliance across the six quality systems during GMP inspections, while understanding the specific expectations associated with pre-approval inspections.


By establishing strong quality systems and clearly defining responsibilities between CDMOs and sponsors, organizations can be better prepared to address regulatory inspections and respond effectively when issues arise.


Read the full article and watch the Ask the Expert discussion:

Audits, Inspections, and CDMOs
 

Link to the Video and Article on Pharmaceutical Technology


Strengthen your inspection readiness with a partner who understands the full regulatory landscape.

RCA supports pharmaceutical and medical device organizations with comprehensive audit preparation, supplier oversight, CDMO qualification, remediation planning, and ongoing quality system optimization. Whether you’re scaling operations or navigating complex regulatory expectations, our experts help you reduce risk and ensure confidence in every inspection.


Take the next step toward true compliance readiness.

Explore our full range of services here: Compliance Assurance Services or connect with our team to discuss your tailored compliance strategy today.
In this episode of the Ask the Expert video series published by Pharmaceutical Technology®, Susan J. Schniepp, Regulatory Compliance Associates, and Siegfried Schmitt, Parexel, give their opinions on why those working in the pharmaceutical industry should lend their voices to draft regulations.


Regulatory agencies, such as FDA and the European Medicines Agencies, and other industry standard setting bodies, such as the International Council for Harmonisation and the US Pharmacopeia, often publish draft versions of the guidance documents they are developing to gather feedback from the public, which includes those scientists and engineers working in the pharmaceutical industry. But who should comment and why?


“You know, there’s an opportunity before regulations get approved for industry to participate and offer their comments on the regulation. This has worked in many cases where, for instance, in the US, way back in 2013, 2014, there was a proposal for metrics. And it was through the commenting period that was open to the public, where industry was able to put forth a case to get the agency to consider some other metrics that we thought were important, that they were not originally in the proposal,” explains Susan J. Schniepp is distinguished fellow at Regulatory Compliance Associates. “So, the way it works is anytime there’s a new regulation or a change to an already existing regulation, the FDA will announce it in what we call the Federal Register. It’s kind of like the daily newspaper for the government. It’s that time when they open it up for comments from industry. And so, it’s very important because you can change the final regulation before it becomes official by putting your comments in through this vehicle.”


“Absolutely anyone can comment,” stresses Siegfried Schmitt, PhD, is vice president, Technical at Parexel. “You don’t have to be the ultimate expert. If you feel you have something to say, absolutely, you can. Now, perhaps it is advisable or sensible to not just comment as an individual but perhaps comment through your company. Perhaps your company as such is interested in providing comments.”


Both Schmitt and Schniepp also suggest commenting through a trade association, such as the Parenteral Drug Association. “I like commenting through industry associations because usually, they gather a group of experts together proposed regulation topic, And you get to hear not only your perspective voiced, but other experts’ perspectives voiced,” Schniepp says.


Developing an Effective Contamination Control Strategy

In this article, RCA expert and Susan J. Schniepp of Regulatory Compliance Associates (RCA), a Nelson Labs company, discuss how contamination control strategies (CCSs) support pharmaceutical manufacturing quality, patient safety, and regulatory compliance. A CCS provides a comprehensive, proactive approach to identifying, assessing, and controlling contamination risks while supporting continuous improvement across manufacturing operations.


The article highlights key components of an effective CCS, including personnel and environmental monitoring, process and equipment validation, cleaning and sanitization, personnel training, and quality risk management. A strong quality culture and scientifically grounded controls are essential to maintaining effective contamination prevention practices. The experts also emphasize the importance of aligning contamination control efforts with applicable regulatory expectations, including EU GMP Annex 1 and relevant FDA and ICH guidance.


Risk assessment is a fundamental part of a successful CCS, helping manufacturers identify potential contamination sources, evaluate their impact, and prioritize preventive measures. By integrating risk-based decision-making, cross-functional collaboration, and ongoing process improvement, organizations can strengthen contamination controls, improve product quality, and protect patient safety.


Read the full article: Developing an Effective Contamination Control Strategy.

In this article published by Pharmaceutical Technology®, Matt Cushing, VP of Quality and Science, Nelson Labs, and Susan J. Schniepp, distinguished fellow at Regulatory Compliance Associates, discuss PDA/ANSI Standard 06-2025: Assessment of Quality Culture Guidance Documents, Models, and Tools, which was published in February 2025.


Q: How can the new document from the Parenteral Drug Association (PDA), PDA/ANSI Standard 06-2025, help manufacturers and laboratories improve their culture and processes?


A: PDA/ANSI Standard 06-2025 is a consolidation of guidance documents, tools, and models to assess and improve quality culture. Companies have probably seen how the state of the quality culture can impact an organization positively or negatively. This document is a great tool to research the key aspects of culture, assess the status of your culture, and create a strategy to maintain and improve the quality culture of your organization.


Watch our FREE on-demand webinar for a deeper dive into the new Quality Standard. Click here to learn more.


Q: What is a quality culture and what does the term refer to?


A: Quality culture is something seen and felt everyday as an employee and as a consumer in our everyday lives. It is hard to put into words, but it is defined well in the PDA/ANSI 06-2025 standard as “the overriding attitude, both expressed and implied, of an organization towards quality” (1).The document breaks it down into two distinct elements: a culture of shared values, beliefs, expectations, and commitment toward quality and a structural/managerial element with defined processes that enhances quality and coordinates individual efforts. It is the cross-functional ownership of quality at all levels, as well as the beliefs and systems that drive our actions each day.


Q: What factors of a quality culture are important in the pharmaceutical industry?


A: The document has so much information on the key factors of a good quality culture. The consistent themes that stand out are:

  • Leadership commitment—quality accountability, recognition, feedback loop, Gemba, visionary, strategic, enablers, respect your team, humility, trust
  • Employee engagement and ownership—mission focused, striving for excellence
  • Consistent standards and expectations —all sites, all teams aligned and striving for the same outcome, courage to do what is right, communicate the behaviors that you expect
  • Collaboration and communication—together, sharing, learning from each other, planning
  • Technical excellence—which was a newer way to group concepts that companies are familiar with, such as mature systems, competence, organizational learning, technology/innovation, and agility.

Q: Can you talk about the application of these factors in a laboratory setting and how it may differ from that of a manufacturer?


A: All of the above attributes apply and are critical for a laboratory’s quality culture, but in the authors’ opinion, some items are even more critical for a testing laboratory to succeed. In a laboratory, the main resource is people, not machines or facilities. Scientific competence and a good understanding of why things are done a certain way is paramount to producing quality results and solving problems for customers. Good leadership and employee engagement can make a huge impact on quality culture and the results achieved for clients.


Q: To what extent is the customer perspective incorporated into quality decision making?


A: The customer perspective must be embedded in strategic planning, quality systems, and day-to-day activities. The customers’ product end-users, along with their regulators, determine what good quality looks like in any particular setting. If contractors are not viewing their work through their clients’ eyes, they will miss the mark on quality. Aligning employees’ responsibilities with customers’ expectations, particularly in the pharmaceutical industry, creates employee engagement as they get excited about what customers are doing. This also creates partnerships with customers that are based on achieving mutual success. What customer doesn’t want to be understood by the provider to whom they are going for a solution?


Reference

1. PDA. PDA/ANSI Standard 06-2025: Assessment of Quality Culture Guidance Documents, Models, and Tools (PDA, February 2025).


Article details

Pharmaceutical Technology®
Vol. 49, No. 5
Page: 34

In this episode of the Ask the Expert video series, Susan J. Schniepp, distinguished fellow at Regulatory Compliance Associates, and Siegfried Schmitt, PhD, vice president, Technical at Parexel, discuss how to address an FDA warning letter citation for a failure to establish a quality control unit.


Link to the Video and Article on Pharmaceutical Technology


After an FDA inspection of a pharmaceutical manufacturing facility, any deviations the inspector might find during the inspection are reported on a Form 483 and sent to the manufacturer. If the company fails to properly respond to the agency, FDA will follow up with a warning letter. A common theme seen on warning letters is that companies fail to properly establish a quality control unit. So how should a company respond if they receive such a citation?


“Well, you see the mere fact that they actually received a warning letter means that their initial response to the observations by the FDA inspectors were considered inadequate or incomplete,” Siegfried Schmitt, vice president of Technical at Parexel, explains. “I think it’s therefore essential to admit that this shortcoming has happened, and so they should make sure that the response they now give to this warning letter is really a complete and convincing one and provides a remediation plan that will be acceptable to the authorities.


In this episode of the Ask the Expert video series, Susan J. Schniepp, distinguished fellow at Regulatory Compliance Associates, and Siegfried Schmitt, PhD, vice president, Technical at Parexel, discuss what manufacturers should do when they receive a warning letter from FDA for an insufficient quality control unit.

Susan J. Schniepp, distinguished fellow at Regulatory Compliance Associates, and Rona LeBlanc-Rivera, PhD, principal consultant, Regulatory Affairs at Regulatory Compliance Associates, answer some questions about FDA’s January 2025 21 CFR 211.110 guidance document.


Q: How does FDA’s guidance, Considerations for Complying with 21 CFR 211.110, impact our current approach in complying with 21 Code of Federal Regulations (CFR) 211.110 (1)?

A: This guidance is currently in draft. However, when finalized, it will describe areas to be considered to ensure batch uniformity and drug product integrity. The guidance also discusses quality considerations for drug products manufactured using advanced in-process manufacturing techniques. Allowing flexibility to use new technological advances to determine drug product integrity and batch uniformity during manufacturing could reduce product reject rates because in-process results could be reported sooner (perhaps in-real time) and manufacturing corrections could be made to ensure the batch integrity. Drug products manufactured using advanced in-process techniques may have non-traditional quality considerations to determine the suitability of the product, and this guidance allows for flexibility and latitude on how to implement these controls effectively. It should be noted that the scope of this guidance pertains to commercial processes and not to products in development. The drug substance/API is also not within the scope of this guidance.


Q: How flexible is FDA in allowing alternative approaches to in-process controls, and what would be the process for gaining approval for such approaches?

A: FDA does allow manufacturers some flexibility to use better and more efficient methods to meet current good manufacturing practice (CGMP) requirements. It is recommended that companies gather the data and seek early FDA feedback regarding the use of alternative approaches for both in-process and quality parameters before submission of an application. Companies trying to introduce novel and new in-process test controls that yield more accurate and real-time measurements will have more success in getting approval by partnering with the FDA early in the process of introducing the new technology to the manufacturing process.


Q: What are FDA’s expectations for in-process material sampling frequency and methodology?

A: In-process sampling would be dictated by the nature of the drug and manufacturing process. It is not always feasible to obtain an in-process sample. In this guidance, FDA advises that innovative technologies may allow in-line, at-line, or on-line measurements in lieu of physical sample removal for testing.


Q: How does this guidance address the role of in-process controls in preventing drug shortages?

A: In the guideline, FDA states that it “supports the adoption of advanced manufacturing as a foundation for improving the overall quality and availability of drug products for patients.” Exploring how to improve manufacturing oversight through the use of the innovative technologies mentioned above could help enhance supply chain stability and eliminate some drug shortages. Advanced in-process and quality parameter controls may help reduce production failures that contribute to these shortages.


Q: If a more efficient in-process control method could reduce shortages but it is not covered in the guidance, how should companies seek FDA approval?

A: For an FDA-approved drug product, the application holder should gather the necessary data and documentation for filing a supplement (i.e., CBE [changes being effected]-30, PAS [prior approval supplement]) to their application regarding the use of alternative approaches. FDA would then review and provide comments on the proposed change and help identifying the best pathway forward for regulatory acceptance of innovative manufacturing techniques.


This new guidance offers flexibility to manufacturers, including contract manufacturing organizations, to introduce new, more efficient methodologies to measure batch uniformity and product integrity required to determine product suitability to ensure continued access to necessary pharmaceutical products. It is up to pharmaceutical industry to work with FDA and determine the best implementation strategy for realizing the benefits of modernizing manufacturing processes.


Reference

1. FDA. Considerations for Complying with 21 CFR 211.110, Draft Guidance (CDER, January 2025).


About the Article


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].