Archives: Published Articles

Navigating the FDA’s Quality Management System Regulation (QMSR)

The FDA’s Quality Management System Regulation (QMSR), effective February 2, 2026, aligns U.S. medical device quality system requirements more closely with ISO 13485:2016. However, the transition involves more than updating documentation and terminology. Manufacturers must address substantive changes to quality processes, risk management, and inspection readiness.


In this article, Jordan Elder, director of regulatory affairs at Regulatory Compliance Associates (RCA), a Nelson Labs company, discusses critical steps for transitioning from the Quality System Regulation (QSR) to the QMSR and maintaining compliance across global markets.


The discussion highlights the importance of conducting formal gap assessments, updating quality system documentation, strengthening risk management, and training personnel. Elder also addresses key areas requiring attention, including CAPA processes, management reviews, internal audits, complaint handling, supplier controls, and design documentation.


The article further explores how QMSR alignment with ISO 13485 can support global regulatory harmonization while emphasizing that additional requirements may still apply in markets such as the EU and Canada. Manufacturers should also ensure their inspection readiness practices reflect FDA’s updated QMSR inspection approach.


By addressing critical gaps and integrating risk-based quality management practices, medical device manufacturers can strengthen compliance and support access to international markets.


Read the full article:
Navigating the FDA’s Quality Management System Regulation (QMSR)


Developing a Strategic Medical Device Testing Plan

A well-designed medical device testing plan is essential for demonstrating that a device is safe, effective, and performs as intended throughout its lifecycle. As discussed by experts from Nelson Labs and Regulatory Compliance Associates (RCA), an effective plan connects design requirements, risk management, applicable standards, and regulatory expectations to the testing needed to support market access. Testing may include biocompatibility, sterilization validation, packaging and shelf-life studies, functional performance, usability, and software verification.


Testing strategies should be tailored to the device’s intended use, materials, manufacturing processes, patient contact, and clinical application. For manufacturers pursuing global commercialization, identifying regional regulatory requirements early can help reduce redundant testing and support broader market access. While a single study may satisfy requirements across multiple regions, differences in regulatory expectations, documentation, and testing standards may require additional evidence.


The article also emphasizes the importance of early planning, scientifically justified sample sizes, and consideration of often-overlooked factors such as transportation conditions, environmental stability, aging, cybersecurity, and human factors. By engaging experienced testing and regulatory partners early, manufacturers can develop a risk-based strategy that generates the appropriate evidence while helping minimize delays, unnecessary costs, and potential redesigns.


Read the full article: Developing a Strategic Medical Device Testing Plan


Investigating Patient Complaints Involving Prefilled Syringes

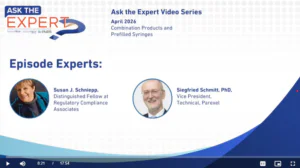
In this episode of Ask the Expert, Susan J. Schniepp of RCA and Siegfried Schmitt of Parexel discuss the challenges of investigating patient complaints involving prefilled syringes (PFS) and combination products. The experts emphasize that marketing authorization holders (MAHs) remain ultimately responsible for product performance, even when purchasing commercial syringe components from external suppliers.


Investigations can be complicated by limited physical evidence, particularly when patients self-administer medications at home. Manufacturers must avoid prematurely attributing device failures to human error and ensure products are suitable for diverse patient populations, including elderly individuals and those with disabilities. Real-world complaints may reveal gaps in usability testing, device performance, or product design.


The experts also highlight the importance of clearly defined quality agreements, effective complaint handling procedures, and collaboration between drug manufacturers and component suppliers. Manufacturers should establish clear investigation responsibilities, adapt complaint processes to evolving distribution models such as direct-to-patient shipping, and ensure corrective actions and effectiveness checks can withstand regulatory scrutiny.


Watch the video: Investigating Patient Complaints Involving Prefilled Syringes


Link to the Video and Article on Pharmaceutical Technology


Data Integrity Hotspots

As artificial intelligence and digital twins become more integrated into pharmaceutical manufacturing and validation, they are creating new opportunities to improve efficiency, strengthen quality oversight, and accelerate decision making. In this second part of a two part interview, Susan Schniepp of Regulatory Compliance Associates discusses how these technologies can transform traditional manufacturing processes while maintaining an essential role for human expertise.


AI can serve as a centralized data aggregator, bringing information from multiple sources together to provide a comprehensive view of a manufacturing batch. By connecting process parameters, deviations, and other relevant information, the technology can help generate a batch dossier and accelerate the product release process. AI can also analyze large volumes of data to identify areas where data integrity risks may exist, allowing quality professionals to address potential vulnerabilities before they become actual compliance issues.


Schniepp emphasizes that these capabilities should complement rather than replace human decision making. A human in the loop remains essential for evaluating the information generated by AI and digital twins and making the final determination on whether a batch is suitable for release. This approach combines the speed and data processing capabilities of emerging technologies with the professional judgment and accountability required in a highly regulated pharmaceutical environment.


Read the full article: Digital Twins and the Future of Pharma Validation, Part 2

Susan Schniepp, Regulatory Compliance Associates Inc., discusses digital twins and AI in pharma, focusing on data integrity, human-in-the-loop roles, and evolving machine learning risks in this publication from PharmTech.


Digital Twins

Link to the Video and Article on Pharmaceutical Technology


In part 1 of a 2-part interview (Link to Part 2), and as a part of PDA Week 2026, Susan Schniepp, Regulatory Compliance Associates Inc., discusses the evolving role of technology in the manufacturing landscape. A central focus of her discussion is the implementation of digital twins, which she describes as systems that mimic live operators. Schniepp explains that when these digital models diverge from physical operations, it prompts corrective actions or new validations to bring the two back into alignment. A primary benefit of this technology is the ability to identify data integrity hotspots, areas where violations might occur due to how documents are handled.


The shift toward AI represents a significant change from traditional paper or middle electronic systems where humans perform every approval. With machine learning, the system can now perform much of the data collection and verification itself, allowing the machine to take on a larger role. However, this shift raises critical questions regarding where human oversight is most efficient. Schniepp suggests that discrepancies between human opinion and machine data should be treated as deviations requiring a formal investigation. She notes, “The traditional model that we have is to do an investigation on it.”


Looking ahead, Schniepp acknowledges the rapid pace of machine learning and its potential to surpass human capability in data processing. She observes, “At some point the machine ultimately is probably more capable at looking at some of these things as it learns, it’s gonna absorb so much more data and information and learn at a faster pace than the human can, that at some point the human’s gonna be, and this is where I get concerned, the human will be out of the loop.” For now, she advocates for human precedence, suggesting that because the technology is so new, humans should investigate the digital twin during conflicts to determine the root cause of any discrepancy.

In this video article published by PharmTech, Susan Schniepp says poor quality in pharma manufacturing is costly but catching defects early can saves millions.

Link to the Video and Article on Pharmaceutical Technology


In part 1 of a 3-part interview, Susan Schniepp, Regulatory Compliance Associates Inc., sat down with PharmTech, during PDA Week 2026, to discuss the critical importance of quality systems in pharmaceutical manufacturing. Drawing on her experience participating in PDA planning committees and her work interfacing with the FDA on 503B compounding, Schniepp makes a compelling financial case for investing in quality at every stage of production.


At the heart of her argument is a straightforward cost escalation model tied to where in the manufacturing process a defective material is caught. Catching a problem at incoming inspection costs virtually nothing, the material is simply returned. But the price of missing that defect climbs steeply from there. As Schniepp explained, “It costs you $10,000 if it makes it on your shelf. It costs you $20,000 if it makes it to your floor.” The numbers continue to rise to $50,000 if the defect enters the batch, $100,000 if it reaches final product, and a staggering $1,000,000 if a full field recall becomes necessary.


Beyond the financial toll, Schniepp emphasizes that recalls carry serious reputational consequences regardless of their severity. “It’s the reputation of the company that’s at stake whenever there’s any kind of recall,” she notes, pointing out that even a cosmetic issue like a smeared label can damage how customers perceive a brand.


Ultimately, Schniepp’s message is clear: a robust quality system is not merely a regulatory obligation, it is a sound business investment. By catching problems early and consistently, companies can avoid the exponentially higher costs that come with letting defects slip through the cracks.