Archives: Published Articles

Good Manufacturing Practice (GMP)


Maintaining good quality control practices throughout the entire manufacturing process requires robust GMP development, a drive toward product and process understanding, and pre-established, comprehensive written procedures that are consistently reviewed and updated.


Good manufacturing practice (GMP) is established by regulators to ensure that pharmaceuticals are safe and effective for the patients that rely on them.


21 CFR Parts 210 and 211


In the United States, requirements governing finished pharmaceutical quality are described in the Current Good Manufacturing Practices (CGMPs) regulations established by FDA and published in the Code of Federal Regulations (Title 21 of the CFR, parts 210–211 for most finished pharmaceuticals).


FDA also publishes guidance to further describe recommended practices for complying with the CGMP regulations. The agency states that “CGMPs provide for systems that assure proper design, monitoring, and control of manufacturing processes and facilities. Adherence to the CGMP regulations assures the identity, strength, quality, and purity of drug products by requiring that manufacturers of medications adequately control manufacturing operations” (1).


Quality Management


In Europe, GMPs are defined by the European Commission in EudraLex–Volume 4–Good Manufacturing Practice (GMP) guidelines (2), which were first published in 1989.


EudraLex states that quality management is “a system of marketing authorizations [that] ensures that all medicinal products are assessed by a competent authority to ensure compliance with contemporary requirements of safety, quality, and efficacy” (2).


GMP Certified 


To harmonize GMPs and other quality requirements worldwide, the International Council for Harmonization (ICH) works with international regulators and industry to develop common guidelines across the industry to ensure consistent quality expectations worldwide.


“At its core, CGMP is a science- and risk-based focus on assuring drug quality. As described in ICH Q10, Pharmaceutical Quality System, an overall attitude to drive meaningful and continuous improvements from the quality unit and other manufacturing employees is essential” (3), FDA told Pharmaceutical Technology.


GMP Manufacturing


The global nature of the pharmaceutical supply chain requires regulatory agencies to inspect and govern GMPs at manufacturing facilities. These inspections sometimes result in actions by regulators (e.g., FDA 483s, warning letters, import bans, and court actions) against companies that fail to follow GMPs.


Common CGMP deficiencies cited by FDA in warning letters sent to pharmaceutical companies inspected during the past year include failures to ensure product sterility, ensure data integrity, create quality control units, and develop and follow written procedures (4–7).


GMP Practices


“Companies should vigilantly encourage GMP practices that reflect the most current and robust methods of processing and control, and with a focus on providing a quality product to US consumers,” says FDA.


“The approach to successful CGMP is not merely a check-box approach where a single obstacle can be identified, nor is it meant to be unchanged throughout the life of a product or a CGMP facility. Using a holistic and quality risk management approach, a manufacturer can select and study specific products and processes with the goal of continual improvement of product quality and quality systems and widespread optimization.”


Quality Culture


So how do companies ensure they are “vigilantly” following GMPs and avoid the wrath of regulators? The answer appears to be found in a company’s “quality culture” and in the development, writing, and following of written procedures. Performing pharmaceutical manufacturing according to GMPs involves a dedication to quality throughout development and manufacturing processes.


“Corporate management plays a vital role in this endeavor by establishing a commitment to quality, which includes providing sufficient resources and oversight for manufacturing operations. There are many elements to successful implementation of CGMPs but measuring and monitoring quality indicators and managing change are key among them,” says FDA.


GMP Certified


According to Susan Schniepp, distinguished fellow at Regulatory Compliance Associates, becoming GMP certified throughout all processes and procedures involves a combination of personnel training, keeping tools and equipment updated, having a strong quality unit, and developing a quality culture at all levels of the company.


“How the company goes about achieving this objective is what is critical. Traditional training may not be enough. There should be constant on-the-job training and oversight on a continual basis. Upgrading tools and equipment, especially computer-driven programs, needs to involve IT departments and adhere to the current data integrity concepts,” says Schniepp.


“No one element will be able to sustain GMPs. All the elements work together to establish a culture where sustaining GMPs is a priority.”


GMP Facility


Chris Moreton of FinnBrit Consulting agrees. “Too often, in my experience, certain factions in an organization think that quality is someone else’s responsibility. There may be an organizational chart that shows where the quality unit sits in an organization, but ‘quality’ (including GMP) is everyone’s responsibility within an organization; from the most senior to the most junior and vice versa.”


Management is key in creating a quality culture, says Moreton. “If the staff see the managers taking an interest and checking on things on a daily basis, the staff will respond, and they will try harder to get things right.”


Regulatory


A commitment to staying current with regulatory expectations is a must, according to Schniepp, and she warns that complacency is the biggest obstacle to maintaining GMPs.


“Once a process or procedure is established and functional, there does not seem to be the impetus to update and revise it as regulatory interpretation and understanding changes. This leaves the process or procedure compliant to outdated standards,” Schnieep says.


GMP Audit


Companies must also learn from prior mistakes, according to Mark Lynch, vice-president of Strategic Compliance. “It’s best to take the lessons learned from product experience and apply them to the culture overall so those lessons only need to be learned once. Those experiences should be continuously applied on the product level, and also used to make improvements to standard operating procedures (SOPs), policies, personnel, and technology.”


Problem solving is another technique that must be honed, according to Lynch. “Companies must also pay continuous attention to problem identification, solution, and improvement. Refinement of problem-solving techniques contributes to organizational learning.”


FDA cGMP


Enforcing quality procedures through a policy that ties failure to follow procedures with grounds for dismissal and using internal audits to detect improper performance are options to ensuring a quality culture, according to Lynch. “It is also important that tools are a regular topic of discussion with operators to capture improvements and assure consistency.


Additionally, sufficient supervisory presence and oversight are key both for monitoring purposes, and to be sure operators can raise questions and get the support they need. Finally, it’s best practice to put a GMP standard reporting mechanism in place that does not require identification, so employees feel comfortable flagging an issue without fear of blame,” he says.


GMP Quality


Companies who do not consistently maintain GMPs may find themselves under additional scrutiny by regulators for repeat offenses. A variety of FDA warning letters have pointed out repeat CGMP violations at companies and/or a particular facility (8, 9).


FDA reports that the agency, “… strives to provide clear guidance to companies proactively and in its enforcement actions. Where repeated violations have been found at the same facility or among different facilities of the same firm, we highlight those violations so that they may be addressed adequately.


Corrective Action


A focus on a commitment to quality is essential to correcting repeated violations, as are adequate corrective actions and procedures. While we generally encourage a focus on overall quality, there are times when we encourage firms to take specific actions, which are included in our warning letters as well as in applicable guidance documents and regulations.”


Repeated offenses may be a failure to look at the big picture and apply solutions across all systems and/or products, according to Schniepp. “Companies think in terms of solving the individual citation but fail to take that answer for change to a global look at fixing other processes and procedures that might be susceptible to the same observation. Tunnel vision when responding to warning letters has a great potential to result in repeat observations,” says Schniepp.


FDA Warning Letter


“Some of these ‘repeat mistakes’ are found to be FDA violations of the same section of the regulations but stem from a different problem. Investigators tend to use repeat findings as a way to point to simplified trends that make one issue appear to be really bad, rather than the complex combination of issues that it truly is,” Lynch says.


“Some companies lack the staff, procedures, capability, and time to fully investigate and solve problems, so they pick something (e.g., a personnel error), close the investigation, and move on to release product, and the same issue reappears because it wasn’t solved.”


Risk Management


Risk management and general cost increases can be another factor to repeated offenses, according to Moreton. “Often cost and/or short-term shareholder interests are used as an excuse to avoid some measures [to long-term change] … People complain that quality costs money, but if they really want to see how expensive things can be, they should try a consent decree.”


SOP Procedures


The lack of written procedures and/or a quality unit is another frequent infraction identified in warning letters. From October 2016 through September 2017, FDA issued more than 400 FDA 483 observations for a lack of written procedures or written procedures that were not fully followed (10).


Quality Program


Written procedures are key to a robust quality program, according to FDA and industry experts. FDA believes that the “most effective quality assurance (and compliance) strategies begin with robust internal procedures to adequately design and maintain a robust operation, and that can quickly identify and correct manufacturing problems when they occur.”


GMP Best Practices


What are best practices for developing written procedures for GMPs? While the agency does not endorse one particular approach to developing written quality procedures, FDA notes that these procedures should “be written to effectively communicate to the users of the procedure.


FDA recommends that the style and format of procedures be accessible to users, as well as ensuring adequate coverage of its purpose. We recommend that the effectiveness of a procedural training program be evaluated to ensure that personnel learn and can follow the procedures as intended.”


GMP Compliance


The trend in a GMP compliance failure to have written procedures is disturbing, according to Schniepp. She suggests that outsourcing quality, especially for start-up companies, may add to this problem. She questions whether outsourcing companies have the processes and procedures in place to handle new products.


Manufacturing Operations


A robust sharing of information between client and contract manufacturing organization is also key, including product and process understanding. “We need to also remember that new products, particularly in the biotech segment of the industry are novel in nature so the old way of doing business may not be applicable. Whatever the reason the industry must focus effort on making sure there are GMP lab processes, procedures, and written instructions in place that support the release of product,” Schniepp stresses.


Having all parties involved in the GMP labeling development of written quality procedures is necessary. “Including everyone affected by the procedure and writing the procedure with their input will result in streamlined and efficient procedures, which will be easier to maintain in the long run,” says Schniepp.


Standard Operating Procedure


Standard operating procedures (SOPs) written by people not familiar with the specific operation can cause disconnects, according to Lynch. “The best way to assure adequate SOPs is to sit down with people most familiar with operations and map out the process steps and handoffs. This can be incorporated graphically using [swim lane diagrams] and similar tools like Visio (Microsoft) and include them as part of the document.”


“In my opinion, in order to ensure that quality procedures are effective, it is necessary to involve those who know the process or operation being documented,” Moreton agrees. “This may mean sitting down with the operator and finding out exactly what is being done and how, not what management thinks should be done and how they think the operation(s) should be carried out.”


Operating Process


According to Schniepp, procedures should be mapped out, committed to paper, reviewed periodically, and updated as necessary. “Companies need to remember that their processes and procedures are not carved in stone and need to be changed to stay compliant with the operations being performed and the current interpretation of regulations,” says Schniepp. New technologies and new product types may necessitate an update to procedures.


Companies should be careful to not try and fit technology or product advances into current procedures but should instead take the time to review their processes and procedures and update them appropriately in responses to these advancements, she says.


FDA Process Validation


Consistent review of GMP procedures is important, especially if a change in equipment, facility, or regulatory requirements has occurred, experts note. “Written procedures should be reviewed and updated as often as needed. However, if a process and procedure is being updated frequently then it probably wasn’t very well written in the first place,” says Schniepp.


“This being said, procedures that are fairly stable should be validated at least every two years to make sure they are still current and reflective of FDA regulatory expectations.” When developing written procedures, says Lynch, “each process and procedure should be tailored to its specific purpose, so they don’t include unnecessary steps that add time to the process without applicable value to the procedure at hand.”


SOP Documents


Companies often lack the documentation that will help operators understand the entire GMP process, says Lynch. “The most important concept to remember when writing procedures is to include the detailed instructions for the operation or processes being defined by that procedure and not include extra explanatory or extraneous information that has no bearing on the operation or process being defined,” agrees Schniepp.


cGMP Consulting


In warning letters, FDA commonly suggests the hiring of a third-party GMP consultant to help companies address their GMP deficiencies. Schniepp says these consultants can provide a “fresh perspective” when resolving GMP issues. “A new and fresh approach is valuable because the consulting firm has no preconceived ideas and can offer new insight to what may seem to be an old and uncorrectable problem,” says Schniepp.


These consultants can be helpful even when not suggested by regulators “because they are looking at the quality with eyes not steeped in the corporate culture,” according to Moreton. Consultants also have knowledge and GMP expertise the company does not have.


“[Consultants] can offer a variety of approaches and suggestions for remediating current problems as well as offering solutions to maintain and improving systems moving forward. One of the most important aspects to consider when hiring a consulting firm is to make sure they not only have the expertise, but they also have the time to devote to fixing the problem,” says Schniepp.


Manufacturing & Industrial Consultant


When hiring a GMP consultant, Moreton suggests companies look at the experience of the contractor as a whole as well as the qualifications of the individual consultants “to ensure there is a good fit with the contractee’s needs.”


Lynch warns, however, that pharmaceutical companies should not rely too much on outside help. “Consultants can help companies get back on track if companies lack the resources internally. However, eventually companies have to sustain compliance themselves. Third-parties should provide expertise in the needed area and technology and have demonstrated success.


Then, they should be able to teach and mentor personnel for improved behaviors and provide flexible models to fit company operations and culture. Usually, this is more than one-time training, but a program of measurement and support over time.”


Article Details
 

gmp pharmaceutical


Pharmaceutical Technology
Vol. 42, No. 7
Pages:18–23, 60
 

To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. 

Insights on driving value from staffing, automation and continuous improvement.


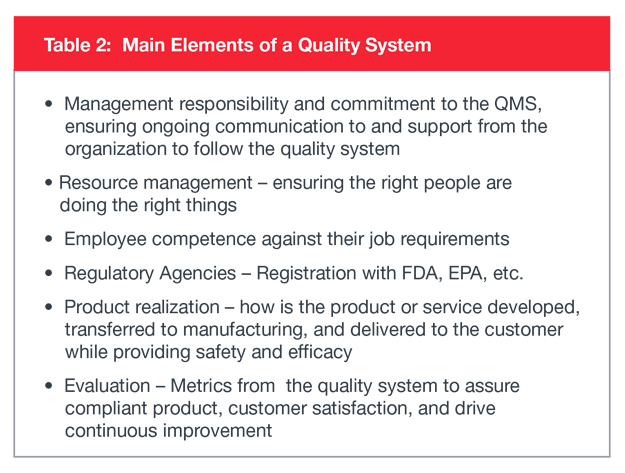
Developing a quality management system is the foundation for ensuring the organization’s products or services are safe, effective and controlled to deliver customer satisfaction. Throughout the organization’s lifecycle, from start-up through maturity, the quality needs of the firm, along with its budget constraints, are continually evolving. Maintaining compliance with regulations while controlling costs represents a challenging balancing act we encounter in our life science consultancy. The most successful firms apply critical assessment of their needs and gaps at present and in the future, and deploy a risk-based approach to their quality system.


Upfront Planning Avoids Costly Surprises


Before establishing a quality system, it’s critical to identify the quality standards, regulations and/or requirements surrounding the company’s products, service and business needs. The checklist in Table 1 provides some common areas to consider.


Gap Analysis


If your organization already has a quality system, perform a gap analysis against your identified standards/requirements per Table 1. If your quality team could benefit from an external assessment, consider hiring an experienced consultant or firm to provide expertise in the gap analysis.


ISO13485


If your organization is or will be manufacturing or distributing internationally, global considerations will add complexity to your quality system. For example, the following considerations can greatly impact the complexity of the quality system; European Union, ISO13485, Canadian Medical Devices Regulations, Brazilian GMP, and Japanese Ordinance #169, and others.


Phased Enhancements: Spread the Spend Over Time


quality management system


Maximizing Value from Staffing


  • Quality System Leader The organization requires a competent and knowledgeable individual as the quality system leader(s) who may implement the QMS, keep the system compliant, and maintain the design history file.
  • Based on the lifecycle of the organization, the quality system leader may be tasked with implementing the QMS or just maintaining the QMS going forward. This could be two different skill sets.
    • Tip: Early stage companies can benefit from outsourcing development of the QMS to highly experienced quality consultants and then transition maintenance of the QMS to less expensive in-house personnel.
  • Skill Assessment By analyzing the skill sets of in-house quality personnel against needs, firms can identify gaps and develop solutions. A few considerations:
    • Tip: Can current staff be trained? Training represents an upfront cost but offers long- term payoff.


Training Programs


  • Utilize assessments and audits to understand the capabilities ceiling of your existing team. If training cannot close the critical gaps, consider whether the needed skill set is temporary or long-term. Temporary gaps can be closed cost effectively through hiring of contractors or DHF consultants to provide specialized knowledge without paying for a full-time employee. Such expertise can be used for:


Maximizing Value from Automation


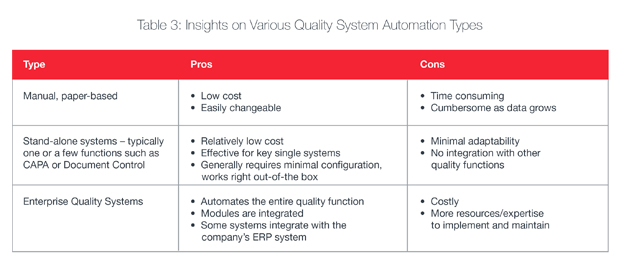
  • Start-up or early stage organizations can benefit from basic manual systems or hybrid systems that automate some of the more labor-intensive quality functions such as document control.
  • Mid-market organizations can benefit from increased automation through hybrid or enterprise QMS systems that address multiple quality needs such as document control, deviations control, non-conformance, equipment calibration, equipment maintenance orders, audit, CAPA, change control, training and functions that control product outputs.
  • Large organizations can benefit from enterprise QMS systems and integrating those systems with their ERP systems for even greater interoperability.


Maximizing Value from Continuous Improvement


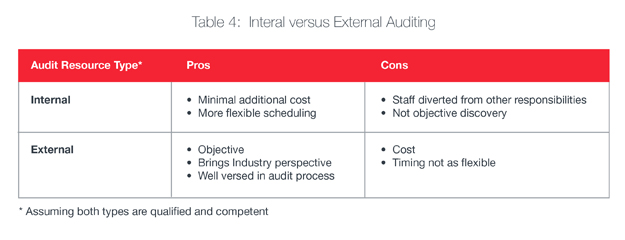
  • Internal and external audits program
  • Complaints
  • Customer feedback
  • CAPA system
  • Product or process non-conformances
  • Processing data
  • Enforcement activity such as a FDA 483 or Warning Letter
  • Costs relative to the budget
  • Concerns about cybersecurity and cost


Budget Maximizing Tips


Travel:

  • Limiting travel by employees and choosing local contractors or consultants to minimize this expense.
  • For organizations with multiple locations, consider leveraging the geography of the company’s quality personnel. For example, if facility A has an out-of-state supplier located near sister facility B, then facility A might be able to tap into the facility B personnel to perform the supplier audit. The travel savings could offset the cost to quality facility B personnel on the QMS audit.

Training:

  • Delaying training can minimize this expense. Although it can be less expensive in the long run to raise the skill set of your employees in areas such as internal audits, some companies trim budgets by foregoing to deferring training.
  • Virtual cGMP training is another possibility to minimize training expense. By training all your employees at once, companies can negotiate reduced training fees and may possibly eliminate department travel by having a trainer come onsite to the company.
  • Delaying supplier audits Sometimes audits can be scheduled to the next quarter or early next year to save expense based on the acceptable risk exposure.
  • Group input: The quality team is the cornerstone of the QMS. Engaging these internal experts can identify creative opportunities to trim expenses while minimizing risk.


Today’s Quality leaders are responsible for managing the organization’s compliance and cost of quality, while driving customer satisfaction and maximizing their department budgets. Careful planning and judicious use of human capital and other resources are critical in deploying a risk-based strategy that delivers organizational results.


To begin the Regulatory Compliance Associates® scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. 


Integrating Data Integrity Into Annual cGMP Training

FDA’s guidance on data integrity reinforces the importance of ensuring personnel understand how to identify and prevent data integrity issues. But organizations do not necessarily need to create an entirely new training program to address these expectations. Instead, data integrity principles can be incorporated into existing cGMP, good documentation practices (GDP), and investigations and CAPA training.


This article explores how companies can use their existing quality systems to strengthen data integrity awareness. Key areas include proper documentation practices, correcting errors, timely signatures, electronic signature controls, password protection, and understanding the responsibilities of operators, verifiers, and reviewers.


The article also highlights the role of investigations, CAPAs, internal audits, and SOPs in maintaining reliable records throughout the manufacturing process. By reinforcing data integrity concepts through routine training and quality processes, organizations can help ensure that both paper and electronic records accurately and reliably support product quality and patient safety.


Read the full article published by PharmTech:

cGMP Training for Data Integrity

Lisa L. Michels, General Counsel and regulatory affairs expert at Regulatory Compliance Associates® Inc., discusses how the global medical device regulatory landscape is impacted by international law.


The global medical device regulatory landscape is constantly evolving as governing bodies and/or agencies worldwide continue their efforts to harmonize and streamline rules. Over the last several years, many of these regulatory bodies have joined forces to ensure consistency in the interpretation, application, and implementation of relevant medtech regulations and globally recognized consensus standards.


Regulatory Submission


While such efforts have considerably improved the product registration process overseas, harmonization in some cases has made it difficult for some device firms to stay abreast of the planet’s evolving regulatory landscape. To remain competitive in this challenging environment, medtech companies stay ahead of the competition by proactively planning for new and/or changing global regulations. This helps agile organizations successfully launch existing, modified, or novel products in new target markets quickly. 


Medical Device Registration


Most companies want their products to be registered in as many global markets as possible, but this lofty goal introduces certain regulatory challenges. From a general business perspective, global product registration in numerous foreign markets obviously makes sense.


Companies possessing the legal authority to sell product(s) in certain regions where competitors are not authorized to do so often distinguishes those organizations as leaders or industry trailblazers. This approach is not always logical or practical, however.


Regulatory Pathway


Product registration should be based on a proactive strategic regulatory analysis. Understanding the available regulatory pathway options can help properly determine whether registration in a particular market. Ultimately, the team is challenges with the regions that make sense for the product and business as a whole.


The most common international medical device registration challenges are primarily based on the following considerations:

  • Understanding and applying country-specific medical device regulations across global markets
  • Timely implementation of country-specific medical device regulations in target markets
  • Costs associated with global product registration(s), which may include (but are not limited to) in-country agent representation, in-country product testing, and in-country collection of clinical data
  • Excessive delays and/or long lead times for regulatory reviews needed for global product registration(s) in target markets


Device Classification


These challenges are much easier to manage when they are properly assessed well before starting the product registration processes in certain global markets. Based on the device classification and type of product, there may be more (or less stringent) requirements that must be achieved in specific countries.


The risk level of the product and the existence of similar approved products in a particular market can complicate the product registration process in certain countries.


Medtech Programs


Many medtech companies typically tend to focus their medical device product registration efforts on seven (7) primary global markets: the United States, Canada, European Union (EU), Australia, Brazil, China, and Japan. Due to ongoing harmonization efforts, these countries are working together to streamline the registration processes and improve their consistency. 


Depending on the company’s overall product launch strategy, device registration in emerging markets has become common in recent years. In certain cases, product registration may be less challenging in these markets because regulations have not yet been fully developed.


Pre-Clinical Testing


The applicable medical device regulations in target markets each pose their own product registration challenges. Therefore, it is imperative that companies plan for all target market requirements early in the product development process—ideally at the idea stage.

Understanding and preparing for the hurdles associated with pre-clinical testing, clinical testing, performance testing, safety testing, labeling, and compliance with harmonized standards, can help companies overcome possible challenges with the product registration process in each potential target market.


Regulatory Compliance


Achieving product registration compliance in each target market may seem like a daunting endeavor, but it’s not truly as complicated as it sounds. There are many similarities between the various regulations, as outlined in the table at right. Although the table represents only a subset of requirements, the country-specific mandates applicable to product type in each of the target markets must be carefully reviewed and assessed.


The most effective method to accomplish this task is to prepare a global regulatory strategy for the product(s), which may ultimately be used as a gauge to determine the likelihood of successfully launching a device in a particular market.


Regulatory Strategy


A global regulatory strategy is a documented analysis that defines the overall business objectives and requirements necessary for foreign product registration. A regulatory plan, on the other hand, is a comprehensive report based on the foundational regulatory strategy that outlines all product- and country-specific registration mandates in particular markets. 


The regulatory strategy and plan may be separate stand-alone documents or combined into a single global strategy and planning file. Regardless of format, however, their purpose is the same—to proactively document and plan for the implementation of applicable regulatory requirements associated with specific products in each target market.


Regulatory Audit


Keep in mind that these regulatory strategy and planning document(s) are not static. They must be updated and revised accordingly when changes are made either to the product itself or to the device’s launch strategy. A revision would be required, for example, if target markets are added or removed from the launch strategy. Any regulatory body will want to understand the risks made to existing products and regulations (particularly when new rules are introduced that can significantly affect or completely alter product registration mandates).


The format of the regulatory strategy and planning document ultimately depends on the company. Some organizations encourage the use of a specific format or template to document the regulatory strategy and/or plan. Other firms are more liberal about format.


Medical Device Regulatory


Fundamentally though, the setup of the regulatory strategy and planning document is not as important as the goal: To initiate and maintain robust, proactive, regulatory planning efforts as early as possible, and to effectively monitor any changes in the scope to avoid unexpected and costly delays. The key elements of a global regulatory strategy and planning document are also outlined in the table. 


Medical Device Global Regulatory Strategy and Planning Document


Detailed Product Summary

  • Detailed description of the product including its accessories, components, and software (if applicable)
  • Describe the product requirements, including technological/functional/performance/clinical requirements of the product (e.g., what the product does and how it works/mode of operation)
  • Indications for use/intended use
  • Target population
  • Labeling requirements
  • All proposed marketing claims and requirements for claim substantiation
  • Risk level/product classification/applicable product codes
  • Predicate(s) and/or similar products on the market in each target country


Governing Regulatory Body and/or Agency in Each Target Market

  • Planned target markets for product launch
  • Applicable laws, regulations, standards, and relevant guidance for the product
  • Analysis of harmonized requirements and standards for the product across all target markets
  • Reimbursement requirements and other country-specific considerations


Proposed Regulatory Pathway and Product Registration Requirements in Each Target Market

  • Regulatory submission/product registration requirements, including all applicable product testing requirements (e.g., pre-clinical testing, animal testing, clinical testing, performance testing, safety testing, etc.)
  • Timeline to obtain clearance/approval
  • Costs associated with global product registration
  • Required resources (internal/external)
  • Pre-market submission/early interaction, discussion, or consultation with governing regulatory agency or authority regarding product and proposed regulatory pathway


General Requirements for Product Registration in Each Target Market

  • Implementation of a Quality Management System (QMS) compliant with applicable regulations and/or standards
  • QMS audit/inspection of manufacturing facilities by the regulatory agency


Country-Specific Requirements for Product Registration in Each Target Market

  • Appointment of an in-country, local agent/representative/sponsor/distributor to manage the product registration on behalf of a foreign manufacturer
  • In-country product testing requirements and product samples
  • In-country clinical trial/data collection requirements
  • Product registries/databases
  • Labeling requirements
  • Language translations for labeling [e.g., instructions for use (IFU), operator or user manual, labels, etc.]


Other information as applicable for the specific type of product


Recommendations for Successful Navigation


The key to successfully navigating through the regulatory challenges of global product registration in the international marketplace starts and ends with diligent and proactive planning.

Product registration efforts should focus on markets where companies can leverage device submission documentation for previously approved applications. If, for example, a company has U.S. Food and Drug Administration clearance and/or CE mark approval for a product, it is often easier to register that exact device in another global market that recognizes the same (or similar) harmonized requirements and standards.


Companies should ensure their products are tested and comply with globally recognized consensus standards. A sampling of medical devices standards recognized in many international markets include:


  • EN 1041:2008—Information supplied by the manufacturer of medical devices
  • EN ISO 13485:2012—Medical devices – Quality management systems – Requirements for regulatory purposes (ISO 13485:2003)
  • EN ISO 14155:2011—Clinical investigation of medical devices for human subjects – Good clinical practice (ISO 14155:2011)
  • EN ISO 14971:2012—Medical devices – Application of risk management to medical devices (ISO 14971:2007, Corrected version 2007-10-01)
  • EN 60601-1:2006—Medical electrical equipment—Part 1: General requirements for basic safety and essential performance
  • EN 60601-1-2:2007—Medical electrical equipment—Part 1-2: General requirements for basic safety and essential performance – Collateral standard: Electromagnetic
  • EN 60601-1-6:2007—Medical electrical equipment—Part 1-6: General requirements for basic safety and essential performance – Collateral Standard: Usability
  • EN 62304:2006—Medical device software – Software life-cycle processes


A comprehensive global regulatory strategy and planning document must be prepared that clearly identifies all county-specific requirements necessary for successful product registration in each target market. In addition, companies should set realistic and attainable goals for timely product launches only in the markets that align with their overall global business strategy. Commercialization will only be successful if the product is a logical fit for the chosen market.


Perhaps most importantly, medtech firms must not register a product in a certain market simply because they can do so. Rather, they should make strategic and informed regulatory decisions about their product launch plans so they can avoid the most common regulatory challenges impacting this highly competitive industry. 


To begin the Regulatory Compliance Associates® scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. 


By Seyed Khorashahi, MSc, and Mark Agostino, MSc


The purpose of this article is to highlight new facets of EU Medical Device Regulation (MDR) in the medical device industry. The article contains references to both MDR legal articles and recommendations that will challenge organizations to take a more holistic viewpoint of their products, resources, and regulatory toolkit to be compliant in the EU.

Introduction

medical device regulationThe application date of 2017/745 MDR1 is 26 May 2021, when it will officially supersede the 93/42/EC Medical Device Directive (MDD) that came into effect in 1993. Medical device companies that market their products in the EU are now responsible for meeting new, comprehensive requirements and compliance expectations during the entire lifecycle of their products.


Every medical device manufacturer, importer, and distributor who wants to continue marketing their product into the EU or initiate business in the EU after 26 May 2021 will be responsible for MDR compliance.2 This is a significant change for many different organizations around the world. The rigor required by EU notified bodies will affect the time and resources companies need to become Medical Device Regulation compliant based on the risk of the device.


Have questions about MDR Implementation? Contact Us Now →


Lifecycle management


Lifecycle management in the medical device industry is constantly evolving because of new legal regulations; the complex risks associated with modern technology; the more advanced levels of oversight for legacy products; and increased exposure of development cycle process gaps. These changes present a challenge for notified bodies to become accredited under the new regulations, resulting in a smaller number of MDR-designated notified bodies.


Consequently, the timelines for conformity assessments of quality system and technical documentation have also increased. This directly challenges organizations to develop more proactive lifecycle strategies when preparing to obtain their CE mark,3 the formal sign of conformity received from a notified body indicating that the device has met both quality system and technical documentation requirements of the EU Medical Device Regulation and can be placed in the EU market.


Product Lifecycle


The foundation of MDR legislation is based on historical product lifecycle issues and quality concerns. These new legislative parameters are designed to drive an increase in both regulatory education and corporate accountability across the entire industry.4


Economic operators in the supply chain become responsible for reporting complaints to the device manufacturer, which includes registering medical devices distributed across their supply chain to healthcare providers. Notified bodies also now have a legal responsibility based on product lifecycle quality control and can be held liable based on the manufacturer device class.


Postmarket surveillance


Postmarket surveillance (PMS), vigilance, and market surveillance are covered in articles 83-100 of the MDR, and address/cover the following:5


  • Postmarket surveillance system of the manufacturer.
  • Postmarket surveillance plan.
  • Periodic safety update report.
  • Reporting of serious incidents and field safety corrective actions.
  • Trend reporting.

PMS and vigilance activities are meant to drive awareness and initiate field corrective actions addressing field-related issues. In addition, these activities help assure sufficient knowledge of an evolving device technology landscape to assess the benefit-risk profile for a medical device.


PMS as described in the articles 83-86 is designed to increase the accountability and reporting visibility of data included in clinical evidence plans and reports. Vigilance, as described in the articles 87-92, tend to be more reactive and deals with reporting of serious incidents and field safety corrective actions.


An effective PMS program provides:


  • Real-world experience using a broad spectrum of physicians and patients, outside the confines of pre- and postmarket trial(s);
  • Early warning signs of problems by continuously and systematically collecting and evaluating data;
  • Incentives for early corrective action, such as initiating corrective and preventive actions or a device recall;
  • Increased compliance with relevant legislation; and
  • Additional value beyond compliance (e.g., usability).


Vigilance


Vigilance defines the type of incidents that medical device companies report, which can affect the long-term risk associated with both the device class and timelines for recertification. Guidance documents from the Medical Device Coordination Group can clarify the interpretation of the new regulation and increase understanding of the planning and resources needed from the manufacturer based on these new risk qualifications.


Corrective Action


Manufacturers will need to be agile enough to react to the data analysis and quickly address necessary corrective actions. Having a cross-functional triage process driven by risk management can help the regulatory team make appropriate risk-based decisions. Through the analysis, the benefit becomes a deeper understanding of periodic safety, complaints, literature, and overall performance of the device.


One should also consider need for oversight of current products already sold in the marketplace. Reporting must be approved by the person responsible for regulatory compliance, and clinical approval relies on the expertise of the individual creating the reporting.


Risk Management


Risk management processes to gather information from the field will help address the severity of new issues or recurrence of existing issues. Manufacturers and organizations will now have accountability to remedy issues and have accountability if recurring or existing issues are not resolved. Senior level executives must drive accountability throughout the organization to increase the level of accountability by the entire regulatory and quality assurance team.


Internal Audit


Many medical device companies are learning as they go while still conducting their necessary daily business. They should also consider how much new accountability is needed across their respective enterprises. There are new educational resources that can help companies understand the regulation and resulting accountability gaps, but a greater level of investigation will be achieved primarily through a detailed internal audit.


Unique Device Identifier (UDI)


The incorporation of unique device identifiers (UDIs) in the Medical Device Regulation will help with the traceability of devices, similar to the US Food and Drug Administration UDIs, and will enhance the effectiveness of PMS and vigilance. The UDI requirements define a more targeted approach to field safety corrective actions and supply chain monitoring, including the prevention of potential counterfeit products. UDI activities comprise of UDI registration, obligation to place UDI on devices, and UDI data submission.


The implementation of MDR UDI has different timelines, depending on the device classifications, but all devices need to complete the UDI registration to obtain a basic UDI device identifier (DI) from one of the issuing entities, such as GS1 or the Health Industry Business Communications Council.


EUDAMED


The basic UDI-DI is the primary identifier referenced in the technical documentation and the main key for records in EUDAMED, the European database for on medical devices. The obligation to place UDI on devices and UDI data submission will be required on 26 May 2021 for Class III and implantable Class IIb devices; on 26 May 2023 for Class IIa and non-implantable Class IIb devices; and 26 May 2025 for all Class I devices.


In addition, there should be a strategy in place for legacy devices if the new elements of MDR regulation are to be completed. The UDI label design for packaging configurations should be considered to ensure compliance with the increased regulatory expectations in the delivery channel.


Harmonization


The potential for harmonization within the medical device industry is increasing as the industry moves closer to a universal label based on requirements from multiple countries and regulatory bodies. Translations may differ slightly between countries because of on language variations, but the device tracking process should be easier once the label is finalized and regulatory approval is given.


Traceability and market surveillance will invite new analyses beyond the design and application of the label.


Regulatory Submission


The requirements notified bodies have to meet under the EU MDR are extensive and are listed on Annex VII of the EU MDR.6 With the new regulation, there will be fewer notified bodies than the number of MDD-designated, so companies should find and engage with their notified bodies as early as possible in the regulatory submission process. MDD legislation allowed for a less stringent certification process for lower-risk classes of devices.


Compliance Standards


With new MDR compliance standards now in place, medical devices that were previously in one class may move into another level of risk. This may increase the accountability needed for select product lines and application use cases.


Be deliberate about reviewing the product portfolio and recognize there may be changes needed in the expertise within the regulatory team. Employees could be cross-trained over time to facilitate compliance within the new MDR process and work with the different notified bodies.


In addition, not all notified bodies have the technical capabilities for performing conformity assessment for all types of devices and technologies, which may also present challenges and slow down the assessment process.7 


Product Classification


If a product’s classification requires involvement of a notified body for conformity assessment, it is worth noting that successful teams often engage early with their notified body to align with their expectations based on the risk level of their products.


Under MDD, notified bodies also had a consultative opportunity to counsel companies, but that will no longer be available to medical device companies. Without this advisory input from the notified body, companies are at risk of legal liabilities as they try to meet the MDR requirements on their own.


Notified Body


It is therefore important they address ways to compensate for no longer having advisory input from the notified body, for example, by working with a consultant who is not associated with the notified bodies. In addition, the regulatory team should identify the professional skill sets needed for engaging with a notified body and develop educational strategies for training employees who will be working with a notified body for the first time.


State-of-the-Art Design


Manufacturers commonly have questions during postmarket surveillance about understanding the difference between what is, and what is not, state-of the-art in design. For the sake of clarity, “state of the art” is intended to define new products that have been developed and approved for sale. This is unique to devices that are already in the field and have some form of legacy CE marking. A new device cannot be considered state of the art until this updated regulatory approval is given.


The term “state of the art” is a widely used term but was specifically defined within the medical device context in ISO/IEC Guide 63:2019,8 which says state of the art is the “developed stage of technical capability at a given time as regards products, processes, and services, based on the relevant consolidated findings of science, technology, and experience.”


Clinical Evaluation


EU MDR mentions the term “state of the art” 12 times but does not define it. MEDDEV’s Clinical Evaluation document9 describes it as the current knowledge/state of the art in the corresponding medical field, such as applicable standards and guidance documents, information relating to the medical condition managed with the device and its natural course, benchmark devices, other devices and medical alternatives available to the target population.


Risk Profile


As new technologies build upon existing platforms, the intent of the term is to ensure that a proposed device technology is considering the benefits and risks of the similar existing devices that are on the market. The objective being driven by EU MDR and notified bodies is that a proposed device technology meets, at a minimum, the current benefit and risk profile or is able to improve upon the existing profile for similar devices.


Software as Medical Device (SaMD)


The technology landscape for medical devices is quickly evolving as new devices such as software as a medical device, wearables, and combination products come to market. The challenge for manufacturers is to be aware of the benefits and risks related to the device technology under development.


Conformity Assessment


Demonstrating an understanding of state of the art and incorporating it into the design and development processes will lead to favorable conformity assessment with a notified body. EU MDR raises the bar on the requirements toward having robust postmarket surveillance, vigilance, and clinical evaluation programs. These programs help manufacturers demonstrate to a notified body that their device technology will go to market with an acceptable benefit-risk profile.


Clinical Evaluation


Clinical evaluation validates the intended use of a medical device and establishes the safety and efficacy in a clinical setting.10 A critical facet of the MDR is to obtain CE marking through the conformity assessment process. There are unique standards and protocols that have been established and must be followed when developing CE marking for regulatory approval.


CEP and CER


The Medical Device Regulation has increased the amount of supporting data medical device companies need to provide to with their submissions for approval of a clinical evaluation, which includes monitoring the performance and intended use of a product based on standards of efficacy. Two of the critical steps of Medical Device Regulation include generating a clinical evaluation plan (CEP) and clinical evaluation report (CER).


Clinical Evaluation Plan (CEP)


The CEP presents the rationale, objectives, design, methodology, monitoring, statistical considerations, organization, and conduct of a clinical investigation. It is the blueprint for demonstrating how the device will meet clinical and performance claims made in the intended purpose throughout its lifecycle. The Medical Device Coordination Group has not published a template for CEP, although it has one for the clinical evaluation assessment report and provides guidance on templates for the PMCF evaluation report and PMCF plan.


The CEP is tightly coupled with PMS, risk management, and usability of the device. As an example, if the usability aspect of a product fails to perform during the clinical investigation as anticipated, then the regulatory or clinical team may incorporate those usability failures as part of the clinical evaluation plan to inform later PMS, risk management, and usability considerations. Other inputs for the CEP include, but are not limited to, sterility and biocompatibility.


Clinical Evaluation Report (CER)


The CEP will also provide a roadmap for creating the CER, which would be submitted to a notified body or competent authority as needed to communicate the overall benefit-risk profile for a medical device. The CER is more detailed and includes:


  • Intended use, device description, device classification, clinical evaluation plan, common specifications, if applicable, applicable standards, product equivalence, and state of the art;
  • Clinical literature review, clinical investigations, and related documentation;
  • PMS, postmarket clinical follow-up (PMCF), and the plan for updates and reporting;
  • Labeling, instructions for use, summary of safety and clinical performance (nonclinical and clinical); and
  • Summary of all available data and conclusions.

The  items provided above incorporate the 4 stages for creating a CER, as outlined by MEDDEV.9 Because the report is submitted to a notified body or competent authority it would require the integration and coordination of a cross-functional regulatory and/or clinical team to capture the full range of necessary information. As such, the report is an important tool for communicating an understanding of the device among regulators.
 

Postmarket Surveillance


The purpose of PMS is to continuously verify the benefits of medical devices throughout the product lifecycle and identify previously unknown risks through observation and analysis of real-world, daily practical usage. If PMS observations suggest changes might be needed in the clinical evaluation plan or report, then PMCF studies must be done to obtain supporting data for updating and revising the CEP/CER to reflect the new findings.


Be proactive in defining how the regulatory/clinical team will monitor uses of the device for both approved and off-label use cases. Also be prepared to proactively integrate your clinical findings and risk management strategies into a cohesive route to regulatory compliance.


Person Responsible for Regulatory Compliance (PRRC)


Medical device manufacturers need to have oversight of product development throughout the lifecycle, lifecycle (from design, manufacturing, postmarket surveillance/vigilance activities, and so on.)


Manufacturers must have at least one person in the company who is a medical device expert and can be designated as the person responsible for regulatory compliance (PRRC). The PRRC ensures the conformity of the device is appropriately checked; the technical documentation and EU declaration of conformity are written up and kept current; and the PMS and vigilance obligations are met. Micro- and small enterprises are not required to have a PRRC but need to have such a person at their disposal.11


Compliance


The PRRC is often designated by a company’s senior management, which underscores the level of responsibility of the position and importance of coordinating collaboration across a number of teams to maintain compliance. The position carries significant legal responsibility for the PRRC because the company could hold them accountable for data quality errors that may lead to noncompliance over time.


As such, companies are required to have liability insurance in case they are sued by EU citizens who might suffer physical, device-related harm.


Summary


Given the extent of the changes under the MDR, organizations are being challenged to take a more holistic viewpoint of their products, resources, and regulatory toolkit to maintain product compliance in the EU. Lifecycle management is just one piece of the puzzle to maintaining regulatory compliance. There are unique approaches to lifecycle management under the new Medical Device Regulation that should be considered.


Postmarket surveillance helps establish a process for identifying and rectifying issues during the course of the product lifecycle. Notified bodies now have a process to absorb the feedback from clinical evaluations to ensure medical devices being designed and manufactured meet the stated intended use. The UDI makes it significantly easier to prioritize what issues need to be addressed and ensure tracking mechanisms are in place so field corrective actions can be accurately executed.


Product Information


That is possible because the UDI has specific product information – lot number, date of manufacturing, expiration date, and so on – which can be used to recall products that are defective. These pieces of information are both human and machine readable (the latter, by barcode or radio-frequency identification), which makes it easier to remove a product from the supply chain.


Quality Management System 


And finally, the quality management system (QMS) is the critical element for the regulatory team to implement and manage a successful PMS strategy.12 This “listening system,” comprised of all the aforementioned elements, provides a company the closed-loop feedback needed from the real-world situations to improve clinical performance.


This helps minimize the cost containment of a product recall by recalling only noncompliant devices based on the specific UDI information, which can help device firms align across the supply chain.


Abbreviations
CEP, clinical evaluation plan; CER, clinical evaluation report; MDD, Medical Device Directive; MDR, [EU] Medical Device Regulation; PMCF, postmarket clinical follow-up; PMS, postmarket surveillance; PRRC, person responsible for regulatory compliance; UDI, unique device identifier; UDI-DI, UDI device identifier.
 
About the authors
Seyed Khorashahi, MSc, is executive vice president of medical devices and chief technical officer at Regulatory Compliance Associates (RCA). 
 
Mark Agostino, MSc, RAPS, is managing director of QARA Biomed and senior medical device good manufacturing practice expert at Redica Systems. His areas of expertise include quality assurance and regulatory affairs for medical device and combination products. He has provided guidance to sponsors and contract service providers on meeting requirement for the design history and technical files for the FDA and under the EU Medical Device Regulation; compliance with ISO 14971:2019; implementing supplier quality processes; conducting internal and external audits; preparing global regulatory submissions; and ensuring compliance to global standards and regulations. Agostino has a master of science degree in biomedical engineering from Worcester Polytechnic Institute, Mass., and an executive MBA from Suffolk University, Boston. He is a member of RAPS and can be reached at [email protected]
 
Citation Khorashahi S, Agostino M. Strategic lifecycle approach to medical device regulation. Regulatory Focus. May 2021. Regulatory Affairs Professionals Society.
 
References
All references were accessed on 24 May 2021. Except for reference 6, 8, and 9, all references are for Regulation (EU) 2017/745 of the European Parliament and of the Council of 5 April 2017, https://eur-lex.europa.eu/legal-content/EN/TXT/PDF/?uri=CELEX:32017R0745&qid=1620164088978&from=EN, with page numbers provided for the specific sections.
 

  1. Reg. 2017/745, Articles 1-4:13-20.
  2. Reg. 2017/745, Articles 1-4.
  3. Reg. 2017/745, Articles 19-20:32.
  4. Reg. 2017/745, Articles 25-34:34-40.
  5. Reg. 2017/745, Articles 83-100:71-82.
  6. Reg. 2017/745, Annex VII:123-139.
  7. Reg. 2017/745,  Articles 51-60:49-55.
  8. International Organization of Standardization/International Electrotechnical Commission. Guide to the development and inclusion of aspects of safety in International Standards for medical devices: Terms and definitions [3.18]. https://www.iso.org/obp/ui/#iso:std:iso-iec:guide:63:ed-3:v1:en Dated 2019. Accessed 24 May 2021.
  9. European Commission. Guidelines on medical devices. Clinical evaluation: A guide for manufacturers and notified bodies [Directives 93/42/EEC and 90/385/EEC (MEDDEV 2.7.1 Rev. 4) https://ec.europa.eu/docsroom/documents/17522/attachments/1/translations/en/renditions/native.
  10. Reg. 2017/745, Articles 61-82:55-71.
  11. Reg. 2017/745, Article 15:28.
  12. Reg. 2017/745, Article 10:23-5.


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].

Regulatory Compliance Associates® (RCA) Distinguished Fellow, Susan Schniepp discusses establishing a quality culture from the top of an organization down.


Quality culture is a patient-focused approach to quality that is proactive not reactive or procedural, according to Rick Friedman of FDA, in a conference presentation. When an organization is focused on quality, it can detect problems earlier. Further, quality culture helps manufacturers experience fewer deviations, costly remediations and protect its brand, according to Friedman.


Highest Quality


Creating a quality culture means an investment in quality including organizational structure and actions. Moreover, the highest quality standards include support for all levels of the company and making business decisions. 


Top Quality


“Companies with good quality cultures usually have transparency and collaboration between departments,” says Susan J. Schniepp, distinguished fellow at Regulatory Compliance Associates.  “This is critical when there are problems that impact manufacturing. In addition, companies with a robust culture usually solve the problem faster, which helps them maintain high productivity rates,” she says.


Quality Department


The quality department in a bio/pharmaceutical company plays a critical, but not exclusive, role when it comes to ensuring that biologics and drugs are safe and effective. The entire organization, however, must be involved in ensuring quality is built-in.


“Let’s clear up one issue right away,” says Schniepp. “When we refer to quality culture, we are referring to the culture of the company and not the culture of the quality department. Having an established quality department does not mean a company has a culture of quality.”


Regulatory Affairs


“In a robust culture, the attention to quality is a responsibility of all employees, not just those who work in the quality department. In a robust quality culture, the quality department is a partner with regulatory affairs and operations, it’s a policing function,” Schniepp stresses.


“Quality culture is an all-pervading attitude and sense of purpose in which quality is foremost in everybody’s activities. A quality department is a part of the organization that monitors quality, (e.g., by end-product testing). However, quality cannot be tested into the product. It is inherent, and has to be built in—by design controls,” agrees Chris Moreton, PhD, vice president of pharmaceutical sciences at Finnbrit Consulting.


Operations Management


According to Friedman, “senior management has a critical ongoing oversight role to ensure suitability of operational design, compliance assurance, and maintenance.” Friedman points to behaviors that can lead to the development of quality culture that includes shared accountability, teamwork, listening, continual improvement, coaching, and integrity.


Production Management


Moreton agrees. “In my opinion, a quality culture means everyone from the CEO down.  This includes production management and the most junior staff put quality (and patient safety) at the top of their agendas.”


Quality requires ‘top-down’ and ‘bottom-up’ approaches. If senior management is not committed/interested, then the junior staff will find it difficult to implement and maintain quality activities.


Quality Assurance


Similarly, if the junior staff are not properly trained and committed to and interested in quality, then quality assurance will not be properly implemented. “I have seen organizations where the commitment to quality was lacking, and there was, in effect, a lack of a quality culture,” says Moreton.


Creating Quality Culture


Creating a quality culture starts with the head of the organization, according to Moreton.


“The CEO has to be committed in both words and deeds. [The CEO] has to hire people who will help shoulder the burden while maintaining the business. This then goes on down through to the most junior staff.”


Performance Feedback


“Performance feedback means employees understand how to enforce the rules. There is a line beyond which an individual must not step, because once they have stepped over it, they will be asked to do so again and again—to thine own self be true!,” he says.


Communication is key, says Schniepp. “Robust quality cultures have data integrity and welcome feedback from all employees regardless of position within the company. When a company has a poor culture, the communication is top down, and employee opinion and input [are] not valued.”


Measuring Quality


How does one take stock in the robustness of quality culture in a bio/pharmaceutical company? “There are studies and data that suggest you can measure the quality culture of an organization by looking at the maturity level of the attributes in their quality management system,” says Schniepp. “Certainly, the premise is if you can measure a company’s attributes (CAPA [corrective action and preventive action] system, communications), you can measure the quality culture of an organization.”


Regulatory agencies may take the organization’s history into consideration when assessing the effectiveness of a company’s quality practices, especially during an inspection. “How many FDA 483 observations have there been, and how efficiently did they respond and rectify the issues? Undeniably, the FDA inspector will take note of how people are working during an inspection.”


Operation Plan


In some ways, it all comes down to the inspection process and the FDA inspector,” says Moreton. “[Regulators] want quality (and patient safety) to be taken seriously, and not simply paid lip-service to. They will look at the operation plan to see who makes the decisions affecting quality.”


Manufacturing Facility


“The general state of the manufacturing facility and the attitude of staff is apparent during an on-site inspections. Consequently, they will analyze how complaints are handled and recall-readiness—how the organization would cope during a recall,” says Moreton.


Data Integrity


“Regulators have linked the lack of a quality culture with data integrity issues. Further, guidance documents released from the Pharmaceutical Inspection Co-operation Scheme, the UK’s Medicines and Healthcare products Regulatory Agency, the World Health Organization, and FDA all stress the link between a robust quality culture and reliable and consistent data.


Employee Feedback


One thing they all stress is an open culture that embraces employee feedback and values employee contributions. “Additionally, senior management can often follow the employee’s lead when establishing a culture of quality,” says Schniepp.


Poor Quality 


Is the investment in creating a company-wide culture of quality worth the time and effort? In fact, it seems the alternative is worse. The repercussions of not having a robust quality culture include higher costs and increased scrutiny by regulators.


FDA 483


“Ultimately, [the impact] is likely to be financial because there will be FDA 483 observations to remedy. After all, product recalls and the possibility of a consent decree takes time and money. (If people think quality is expensive, try a long FDA 483 list, a product recall, or a consent decree). The company’s stock price will also likely take a hit,” stresses Moreton.


“Companies that lack a good quality culture usually have high deviation and FDA investigation rates. For the same reason, this impacts their ability to provide safe and effective medicine to patients. It also costs them more money to perform the investigations making them less efficient,” agrees Schniepp. “In the end, the safety of the patient is what matters most.”


About the author

Susan Haigney is the managing editor of Pharmaceutical Technology.


Article details

RCA


Pharmaceutical Technology
Volume 46, Number 5
Pages: 44–45


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].