Archives: Published Articles

When it comes to pharmaceutical manufacturing, ensuring a strong quality culture is one way to avoid recurring GMP issues, such as data integrity. Susan Schniepp, Distinguished Fellow at Regulatory Compliance Associates, and 2022 PDA Board of Directors Chair outlined the evolution of current quality culture initiatives and showcased some tools to help organizations build quality culture into GMP manufacturing operations during the Nov. 11, 2021, Redica Systems Quality Week webinar, “Past, Present, and Future of Quality Culture.”


“I think that is what we see sometimes in our industry,” she said. “We do not remember how we got some of the regulations that exist today, and we repeat our past mistakes.”


From Quality Metrics to Quality Culture


Schniepp began her presentation with a look at how quality culture evolved from the early 2010s discussion on quality metrics. Key points included the following:


  • On Oct. 5, 2005, CDER Director Janet Woodcock directed pharma manufacturers to build “a maximally efficient, agile, flexible pharmaceutical manufacturing sector that reliably produces high quality drug products without extensive regulatory oversight.” This set in motion CDER’s emphasis on quality in manufacturing.
  • Then, in 2012, the U.S. Congress passed the FDA Safety and Innovation Act (FDASIA). Title VII of the Act allows for risk-based inspection of manufacturing sites, including requesting records in advance, or in lieu of an inspection. Additionally, the Act requires the FDA to provide Congress with an annual report on the state of drug shortages.
  • Around the same time that FDASIA became effective, drug shortages became a critical concern for the agency. Schniepp pointed to aging facilities as one reason for shortages. “Tangential to metrics and quality culture, we had aging facilities, i.e., facilities producing many of the drugs showing up on the drug shortage list were degrading and not being maintained.” Quality issues at these sites had become apparent and potentially causing shortages.
  • In 2014, FDA proposed implementation of a quality metrics system to encourage adoption of quality-supporting practices.


GMP Manufacturing & Measuring Quality Culture


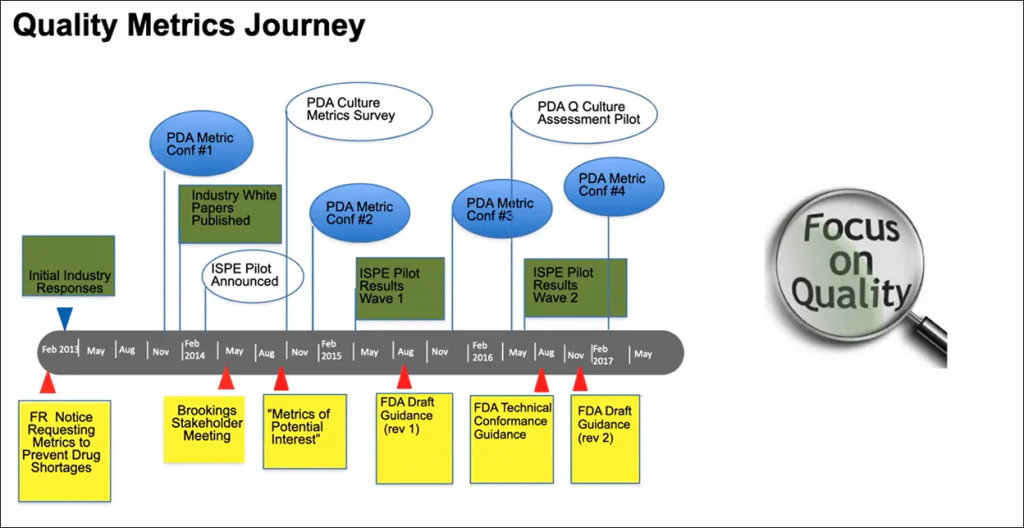
The quality metrics push resulted in a series of conferences by ISPE, PDA, and the Brookings Institution (Figure 1 below).


“One of the elements that came out of all of these discussions was the fact that you could request any metrics that you wanted. What FDA proposed initially was rejection rate on batches, CAPA rates, etc., but the conversation became that without the culture of the organization being assessed, you could not really count on the data. So, that is how [quality metrics and quality culture] tie together,” Schniepp explained.


Quality Metrics


“You needed to have a functioning, efficient, quality metric system that gives you reliable data. And you have to count on the integrity of the culture of an organization in order to have confidence in the data that it sends you. Off of that, the discussion turned to, ‘well, how do you measure quality culture?’”


GMP Manufacturing & Quality Culture Behavior


Schniepp was part of a PDA team studying the relationship between quality culture behavior and mature quality attributes to see if there was a set of mature quality attributes that could serve as a surrogate for quality culture behaviors (ISPE has also done work in this area). 


“’Mature quality attribute’” refers to “those things in your quality management system (QMS) such as your CAPA program, your change control, your investigations procedures, how you report deviations, how you record batch rejection rates, i.e., all of those things are in your quality culture or in your QMS.”


Quality Attributes


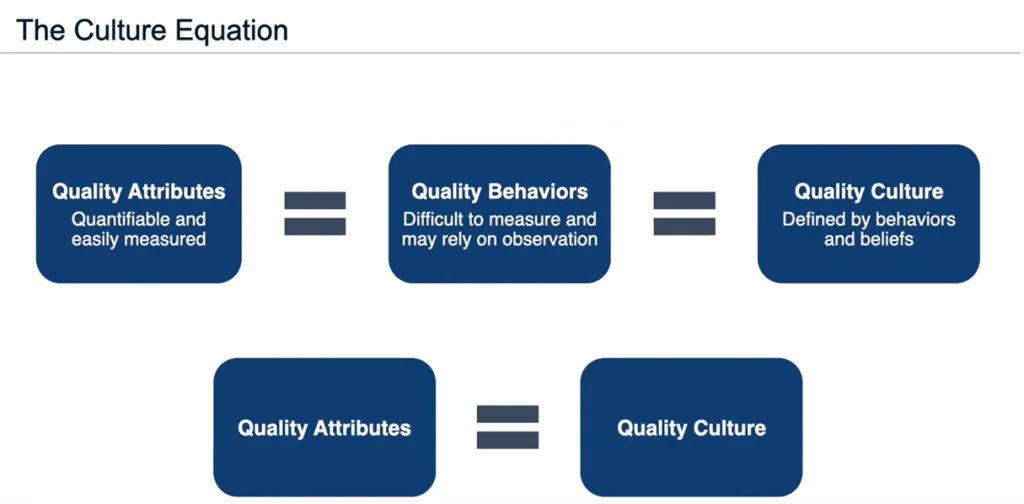
But her team wanted to know if mature quality attributes related to the behaviors of employees and management correlated with company culture, such as do people feel comfortable speaking up, do they talk to management about their concerns, do they understand their jobs, etc. The team’s research resulted in what Schniepp refers to as “the Culture Equation” (Figure 2).
 

regulatory compliance
Figure 2


“The foundation of this is quality attributes that are quantifiable and easily measured, provided they equal quality behaviors,” she said. “In other words, you have attributes reflected in your behaviors that then define your quality culture. So, if quality cultures equal quality behaviors equals quality culture, your quality attributes equal quality behaviors equal quality culture. If you can measure your quality attributes, you can define your quality culture.”


Operational Excellence


By surveying PDA’s membership, her team gathered enough data to conduct statistical analysis and was able to determine there is a relationship between quality culture behavior and quality attributes. They then worked with another team from the University of Saint Gallen’s operational excellence program. That team also found a correlation confirming “quality attributes equal quality behaviors equal quality culture.” 


The PDA and St. Gallen research resulted in the PDA Quality Culture Guided Assessment Tool, addressing five key areas in which companies should focus in order to build quality culture: Employee Ownership and Engagement, Continuous Improvement, Technical Excellence, Leadership Commitment, and Communication and Collaboration. This tool is currently under development as a PDA standard.


Quality Metrics, Data Integrity & Quality Culture
 

regulatory compliance

Figure 1


Data integrity is increasingly connected with quality culture. In the PIC/S guidance, Good Practices For Data Management And Integrity in Regulated GMP/GDP Environments, “they have defined quality culture, and they have given us a little bit of information about how management can create a work environment—a culture—that is transparent and open.” 


Schniepp also pointed to three other documents:

  • UK MHRA GxP Data Integrity Definitions and Guidance for Industry
  • WHO Guidance on Good Data and Record Management Practices
  • FDA guidance Data Integrity and Compliance With Drug CGMP Questions and Answers


“In their regulations, regulators are starting to define and link culture to data integrity violations,” she explained.


Data Integrity


She then provided a real-life example of a data integrity violation caused by a company’s culture.  


“This one is kind of an interesting one because it was really an attempt by management to encourage employees to do a good job, but what they did created integrity violations.”


A contract manufacturing organization (CMO) set a GMP manufacturing goal to release batch records to clients within 30 days of manufacturing the product. To encourage this, management dangled the carrot of a pizza party at the end of the year if this goal was met. In order to meet this goal, employees released batch records to clients even if investigations had not uncovered the root cause. Investigations would be prematurely closed and then reopened after the batch record had been sent to the client. 


“That, in and of itself, is a data integrity nightmare,” Schniepp said. “Can you imagine explaining in an audit by a regulator, ‘well, we wanted a pizza party?’”


GMP Manufacturing & Organizational Culture


Regarding deviation investigations, she told another story of a company for whom she worked that involved repeat observations. When reviewing deviations, she found that one operator had 30 deviations tied to him involving a specific product. This operator failed to sign the batch record during a specific point in time in manufacturing. His managers kept retraining him. 


[Author’s Note: Additional information about deviation investigations can be found in the articles, “GMP Inspection Case Study Focuses on Inadequate Deviation Investigations” and “How to Avoid Three Common Deviation Investigation Pitfalls.”]


Batch Records


In talking with the operator, Schniepp learned that in order for the operator to sign the batch record (according to GMP regulations at that particular point in manufacturing), the operator had to leave the product alone in the equipment, de-gown, sign the batch record, re-gown, and return, all the while hoping the product was okay. Out of concern for the product, the operator chose to stay behind and try to remember to sign the batch record once he was done.


“It was a poorly designed batch record process,” she said. But the operator did not feel listened to when he brought his concerns to management.


GMP Labeling


Schniepp then pointed out that other employees worked on the product under the same cGMP constraints. So, what were these other employees doing? How many other data integrity violations occurred? Was GMP labelling reviewed accurately?

“Nobody else was bringing it up. If you do not have that open culture, if you are not encouraging people to speak up and you are not listening to them, you do not have a good quality culture.”


The Future is GMP Manufacturing Quality


For further evidence of the importance of GMP certification and quality culture, Schniepp reviewed a 483 observation issued in 2019 with numerous examples of data mismanagement. The company in question then responded to FDA with a letter promising to ensure a robust quality culture going forward. [Author’s Note: For more about this GMP compliance incident, read the first case study in the article, “Data Integrity Concerns Discovered in Gene Therapy Product Submissions,” by Jerry Chapman.]


FDA 483


Schniepp believes this 483 observation shows FDA’s intent to heavily focus more on good manufacturing practices in future inspections. This means companies must develop or enhance their quality culture when thinking about how to prepare for upcoming inspections. 


“I believe we will see more of this,” she said. “More companies will be looking at their GMP manufacturing and trying to improve the Quality Culture.”


Otherwise, as she explained at the beginning of her presentation, companies who remain in the past when it comes to cGMP certified practices that lead to quality culture face the prospect of 483 observations, Warning Letters, and other negative outcomes.
 

To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].

Susan J. Schniepp, Distinguished Fellow at Regulatory Compliance Associates, answers some commonly asked questions about aseptic processing and environmental controls.


Q. What is aseptic processing?


A. Aseptic processing is a manufacturing method that can produce product that is absent of bacteria without subjecting the product to terminal sterilization processes.


Aseptic Manufacturing


Many products degrade and become ineffective when subjected to the harsh conditions of terminal sterilization. Aseptic manufacturing allows these products to be produced in a sterile environment, allowing them to maintain their effectiveness while being safe to inject into patients.


Q. What is the difference between aseptic processing and terminal sterilization?


A. The major difference between aseptic processing and terminal sterilization is when the sterilization step occurs in the process. In terminal sterilization, the sterilization is performed after the API, excipients, containers, and stoppers have been assembled.


Sterilization


The final assembled product is then subjected to high heat and/or radiation that renders the product sterile. Terminal sterilization processes are harsh and can have negative effects on product efficacy.


Aseptic Container


For products that can’t withstand terminal sterilization, drug manufacturers employ aseptic manufacturing. The aseptic manufacturing process requires the drug product and any excipients, the container, and the stoppers to be individually sterilized.


Aseptic Clean Room


All must be done before being introduced into the cleanroom or sterile manufacturing core where the final product is manufactured. Additionally, this highly controlled environment is constantly monitored for air quality and potential microbial ingress.


Q. Why must manufacturers establish environmental controls for aseptic processes?


A. Let’s be clear, all drug manufacturing, including solid oral dosage form and terminal sterilization manufacturing are required to have established environmental controls.


Proper Aseptic Technique


This requirement is addressed in global current good manufacturing practices (cGMPs). The purpose of these controls is to prevent product contamination due to unsanitary conditions.


Environmental Controls


The environmental controls to be monitored include, but are not necessarily limited to, air quality including particulate matter, ventilation, temperature, humidity, air pressure, and microbial contamination.


The environment controls have, as expected, stricter limits for aseptic processing manufacturers due to the nature of their business.


Q. What does PUPSIT stand for, and why is it required?


A. PUPSIT is a term used in aseptic processing operations, and it stands for pre-use post-sterilization integrity testing. This testing is performed on the sterilizing filter after it is installed before product manufacturing. Follow up testing occurs again after the product manufacturing is completed.


Filtration Sterilization


The purpose of the pre-test is to ensure that the sterilization and installation process has not damaged your filter prior to the filtration of your product. The purpose of the post-test is to demonstrate that the filter remained intact and undamaged during actual product filtration.


An article published by the Parenteral Drug Association (PDA) states:


“Since 1998, the EU Guidelines to Good Manufacturing Practice: Medicinal Products for Human and Veterinary Use, Annex 1 (Manufacture of Sterile Medicinal Products) or ‘Annex 1’ has contained the requirement for verifying the integrity of a sterilizing grade filter before use and after its sterilization.


The requirement remained in the 2008 revision and in the 2017 draft revision to Annex 1. While not a requirement by the U.S. FDA, EMA inspectors and some PIC/S inspectors have been increasingly expressing expectations for companies to employ this testing procedure”.


The article goes on to explain the concerns that led to the PUPSIT requirement:


“Concerns have been raised that a sterilizing filter could develop certain flaws that would allow microbiological contamination to pass during filtration. The key is that flaws may be blocked or clogged by fluid contaminants or components during the filtration process and remain undiscovered during post-use integrity test. This phenomenon is sometimes referred to as ‘filter flaw masking’”.

Additionally, the article explores the rationale for not employing PUPSIT because of:


“the contamination/product deterioration risk associated with performing PUPSIT may greatly outweigh the risk of product contamination as a result of the masking effect.”


Sterile Filtration


To test a filter during sterile filtration, the sterile filtrate side must be under atmospheric pressure. This requires a fluid pathway to remove any wetting agent.


“The exposure of the downstream portions of the sterile product transport line poses a risk to maintaining the sterility of the filtered product. This, along with other risks, is greater than the remote likelihood of microbiological contamination from a flaw which can be masked during use of the filter that is not detected afterwards.”


PUPSIT


Further, the PUPSIT concept is still actively being debated. The best way to address the use/non-use of PUPSIT in your organization is to make sure you have an appropriate risk assessment in place defending your position.


Q. What makes aseptic drug manufacturing so challenging?


A. Aseptic manufacturing requires highly trained and experienced people to carry out the operations. Even when not in use, special equipment and cleaning procedures must be part of the constant environmental monitoring. Finally, the risk to the product and patients is significant when the aseptic process is compromised.


Article Details


regulatory compliance


Pharmaceutical Technology
Volume 45, Number 10
Pages: 58, 57


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].

Q. I’m a new employee for a start-up company and have been tasked with writing the quality manual for my company. I have searched but have been unable to find regulations that define what is contained in this document. Can you help?


A. The short answer is there is little guidance on what the actual content of a quality manual should contain. It is clear, when reviewing the regulations, there is an intent to have documentation regarding a company’s overall quality approach.


Quality System


In Europe, EudraLex Volume 4, Chapter 1, titled Pharmaceutical Quality System, section 1.7, states:


“the Pharmaceutical Quality System should be defined and documented. A Quality Manual or equivalent documentation should be established and should contain a description of the Quality Management System including management responsibilities”


The United States Code of Federal Regulations (CFR) doesn’t mention a quality manual but does mention a need for a quality policy in the medical device regulations. US 21 CFR 820.3 states:


“Quality policy means the overall intentions and direction of an organization with respect to quality, as established by management with executive responsibility” .


And in section 21 CFR 820.20, the regulation states a,


“Quality policy. Management with executive responsibility shall establish its policy and objectives for, and commitment to, quality. Management with executive responsibility shall ensure that the quality policy is understood, implemented, and maintained at all levels of the organization”.


Quality Manual Examples


The best quality manual examples source for what you should include in your quality manual comes from the International Council for Harmonisation (ICH) in the ICH Q10 document titled Pharmaceutical Quality System. Section 1.8 of this document states:


“A Quality Manual or equivalent documentation approach should be established and should contain the description of the pharmaceutical quality system. The description should include; (a) The quality policy (see section III). (b) The scope of the pharmaceutical quality system. (c) Identification of the pharmaceutical quality system processes, as well as their sequences, linkages, and interdependencies. Process maps and flow charts can be useful tools to facilitate depicting pharmaceutical quality system processes in a visual manner. (d) Management responsibilities within the pharmaceutical quality system (see section III).”


Quality Assurance


Just to be clear, the quality assurance manual applies to the entire company and not just the quality department, similar to how the term quality culture is not referring to the culture of the quality department but the entire company’s approach to culture.


Because the quality manual governs the entire company, it is advisable that it be constructed with teams who have access to the various supplier quality manual data. This collaboration helps ensure the overall purpose and scope of the document is representative of the company’s business.


Purpose Statement


A simple purpose statement might say something to the effect, “The quality system described is intended to ensure the facility is compliant with [list regulations] and serves as the basis of the Quality Systems that apply to [list company name], products, and current good manufacturing practice (CGMP) activities.”


The scope statement can be just as simple and state something to the effect that the quality systems described apply to all products, personnel, and manufacturing operations to ensure products meet the requirements for safety, identity, strength, and purity.


FDA Inspection


Another section to consider adding for FDA inspection purposes is a section of definitions. It is my experience that each company/facility utilizes their own acronyms during an internal audit. Any special acronyms being used at the facility should be defined in the definitions section along with common acronyms such as CGMP, FDA, EU, SOP, etc. Having a definition section helps everyone (employees, auditors, clients, regulators) understand the overall quality system as it applies to your facility.


Management Responsibilities


The next section I would recommend being included is an organizational chart. The organizational chart should demonstrate the reporting hierarchy of the company to demonstrate the independence of the quality organization.

One of the most critical sections to include in your quality control manual, in my opinion, is the section on management responsibilities. In this section, you should introduce the company’s management review responsibilities as well as the senior management titles that are responsible for overseeing the pharmaceutical quality system and the management review meetings.


Quality Management System


The next critical section to include is an overall description of the quality management system in effect at your company. In this section, you can describe your company’s approach to all the applicable systems utilized by your company for planning, documenting, implementing, and monitoring the activities that potentially impact product quality and patient safety.


These processes included but are not limited to quality by design, quality risk management, corrective and preventive actions (CAPA), change control management, management of materials, and maintenance of facilities. In describing the quality management system, you should discuss how it relates to manufacturing, facilities and equipment, materials, packaging and labeling, and laboratory controls. If you outsource an activity, you should note that the activity is outsourced and your oversight responsibilities.


Approval Process


The last element of quality assurance for your quality manual is its approval process. In my opinion, the quality manual should be agreed to and signed off by representatives from the various departments it governs. The final step in the process is to ensure that everyone, from senior leadership on down, is trained and understands the quality manual and how it relates to their job responsibility.


If you follow this simple approach to constructing a quality manual you will have an easy-to-follow, comprehensive document that communicates the company’s commitment to manufacturing safe and effective pharmaceutical products.


RCA


Pharmaceutical Technology
Vol. 46, No. 2
Pages: 50, 49


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].

Developing Effective CAPA Plans After an FDA 483

This article addresses best practices for developing a corrective and preventive action (CAPA) plan following an FDA 483. A well-designed CAPA system should identify the root cause of nonconformances, system failures, or process problems, implement appropriate corrections and corrective actions, and establish measures to prevent recurrence. The resulting investigation and actions should be thoroughly documented and support both regulatory compliance and continuous improvement.


The article emphasizes the importance of timely and complete responses to FDA observations. Each observation should be addressed as a separate CAPA, with the response documenting the investigation, root cause, containment measures, corrections, planned corrective actions, and expected completion dates. Actions already completed at the time of submission should also be supported with appropriate documentation.


FDA may continue to evaluate a company’s response after the initial submission, and a lack of immediate feedback should not be interpreted as acceptance. Companies should follow up with evidence demonstrating completed corrective actions and verification of effectiveness. If FDA determines that a CAPA response is incomplete, additional actions may be requested. An objective review by an experienced third party can help ensure that the response addresses the cited observations comprehensively and that proposed actions are practical and appropriately implemented.


Read the full article: Responding to FDA CAPA Requests

Quality Culture and Data Integrity

data integrity

This article examines the growing connection between an organization’s quality culture and the integrity of the data generated to support pharmaceutical product quality. Regulatory authorities have increasingly emphasized that reliable, accurate data depends not only on data integrity tools and procedures, but also on an organizational culture that encourages transparency, accountability, and the open reporting of failures and mistakes.


The article reviews the evolution of regulatory expectations surrounding data integrity, including FDA’s quality metrics initiative and guidance from organizations such as FDA, MHRA, WHO, and PIC/S. Common data integrity observations have included insufficient training, system failures, inadequate procedures, repeated human error, inappropriate system configuration, and intentional falsification. Across regulatory guidance, management leadership and organizational behavior are identified as important factors in maintaining complete, consistent, and accurate data.


Organizations can assess quality culture by examining both quality system attributes and the behaviors that support them. Areas such as deviation reporting, change control, CAPA, complaints, and environmental monitoring can be evaluated alongside transparency, communication, employee engagement, and cross-functional collaboration. By proactively assessing quality culture and addressing weaknesses, companies can better identify data integrity risks and support compliance with ALCOA principles before regulatory inspections.


Read the full article: Quality Culture and Data Integrity

Q: How much CAPA data do I need to support opening a corrective and preventive action investigation?


A: This is an interesting question because it raises the issue of whether a company is overusing or underusing the CAPA system. One data point is enough to open a CAPA depending on whether that data point is a significant occurrence. On the other hand, multiple similar deviations may be overlooked because they are deemed minor with little impact on operations. The most effective way to determine whether you should open a CAPA is to perform a risk assessment of the data.


Preventative Action


regulatory compliance

A single significant deviation, such as putting the wrong label on the product, should justify opening a CAPA. This type of production error carries a high risk and should consider preventative action Sops. If the mislabeled product is released with the incorrect label, it creates a serious risk to patient safety. The CAPA is necessary because it will enable the company to determine the appropriate actions needed to prevent the issue for recurring.


CAPA Manufacturing


In addition, the CAPA will prompt the company to review quality data across manufacturing lines and manufacturing facilities. This type of CAPA quality corrective solution to prevent recurrence can be implemented across the company. An example of a single CAPA records data point that does not necessarily need a CAPA opened might be a field complaint of a bottle of tablets that contained 99 instead of 100 tablets.


CAPA Quality


This occurrence should be investigated, but because it is low risk and more of an inconvenience to the patient, it may not require a CAPA. Opening a CAPA for this one event could be considered overusing the CAPA system. That being said, if the complaint department tracks these data and finds a significant upward trend of short count bottles over a short period of time, a CAPA may be warranted.


CAPA Corrective Action


Underusing the CAPA system is also a concern. Not considering corrective action when faced with CAPA records that suggests there is multiple occurrences of the same/similar deviation prevents the company from continuously improving processes. Let’s suppose that in reviewing deviations you notice that a few manufacturing employees have multiple deviations for the same minor issue in the same batch record over a short period of time and that the deviation has been determined to be human error and only retraining of the operators was performed.


CAPA Quality Assurance


In this scenario, it would be prudent to open a CAPA for several quality assurance reasons. First, human error is rarely the cause of a deviation. Second, obviously the retraining to prevent the error is not effective because the error keeps occurring. Third, the data are telling you that you may have an issue in how effective your process for investigating deviations is being implemented.


CAPA Audit


Conclusion


There is no hard and fast rule on how much data you need to open a CAPA. It could be one data point or several data points. The best tools to use to determine whether a CAPA is needed is a risk assessment of the data and common sense.


RCA


Pharmaceutical Technology

Vol. 47, No. 4
Page: 50


To begin the Regulatory Compliance Associates scoping process today, please enter your information in the blue form below and click the submit button at the bottom of the webpage. You may also email us at [email protected].